Journal of Chemical Theory and Computation 2026
From planar bilayers to vesicles, diverse lipid membrane descriptors are written in LiMBS, a common representation language. With the encoding of the membrane composition, leaflet organization, geometry, and molecular resolution in a human- and machine-readable form, LiMBS supports reproducibility, interoperability, and data-driven computational membrane research.
Langmuir 2026
Using MARTINI coarse-grained molecular dynamics simulations, this study investigated how S-palmitoylation influences claudin-lipid interactions and membrane organization. Palmitoylation promotes claudin localization in ordered membrane domains, increases local cholesterol enrichment, and reduces interactions with disordered lipids. Importantly, the depth of the palmitoylation site determines membrane preference, with shallow sites favoring ordered domains and deeper sites favoring disordered environments. These findings highlight palmitoylation depth as a key structural factor regulating claudin organization in cell membranes.
The Journal of Physical Chemistry B 2026
Post-translational modifications (PTMs) such as phosphorylation, acetylation, and methylation alter protein function by changing local hydropathy and protein interactions. Using the PARCH hydropathy scale, this study shows that phosphorylation strongly increases hydrophilicity, while acetylation and methylation produce more context-dependent effects on the local protein environment.
Langmuir 2025
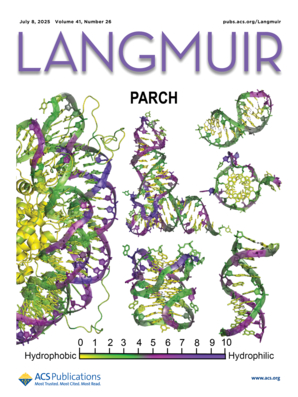
This study extends the PARCH hydropathy scale to Deoxyribonucleic Acid, Ribonucleic Acid, and nucleic acid–protein complexes to quantify residue-level hydropathy and understand their structural interactions. The analysis shows that nucleic acid backbones are far more hydrophilic than bases, RNA is generally more hydrophilic than DNA due to greater structural complexity, and PARCH can reveal key hydrophilic and hydrophobic interaction patterns in nucleic acid–protein binding.
The Journal of Chemical Theory and Computation 2024
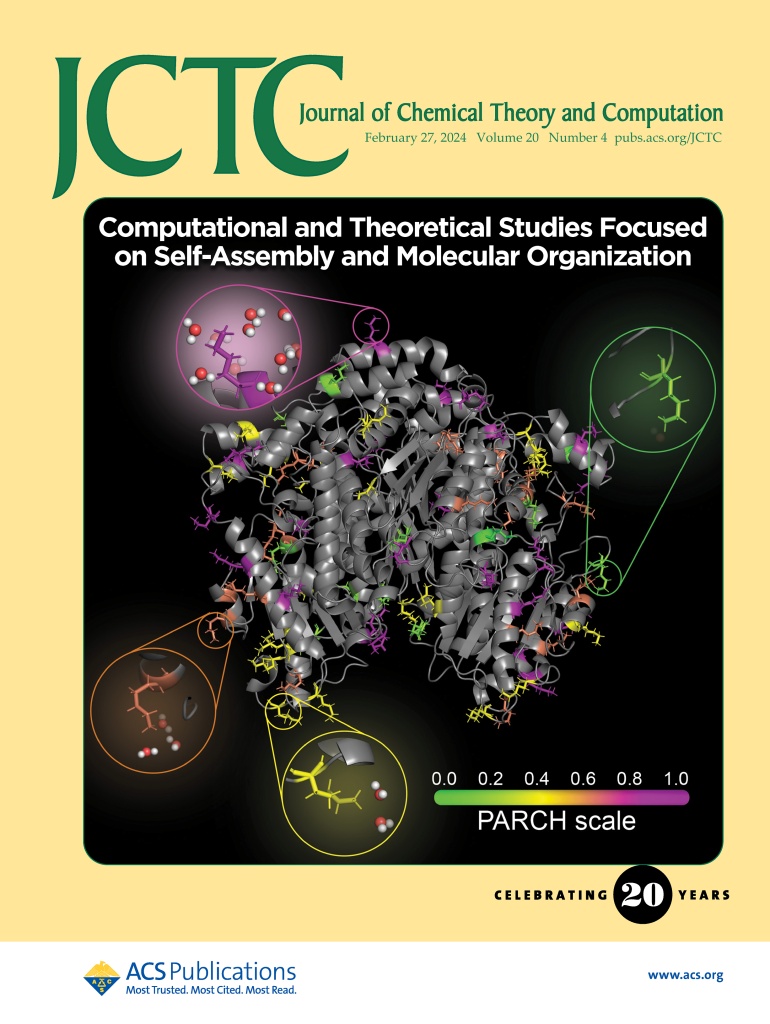
The PARCH scale is a tool for calculating the hydropathy of amino acid residues as a function of a protein’s nanoscale topography, such as bumps, crevices, cavities, clefts, pockets, and channels. This computationally inexpensive method can compare hydropathies of the different protein surfaces and quantify the effect of point mutations.
BioMacromolecules 2023
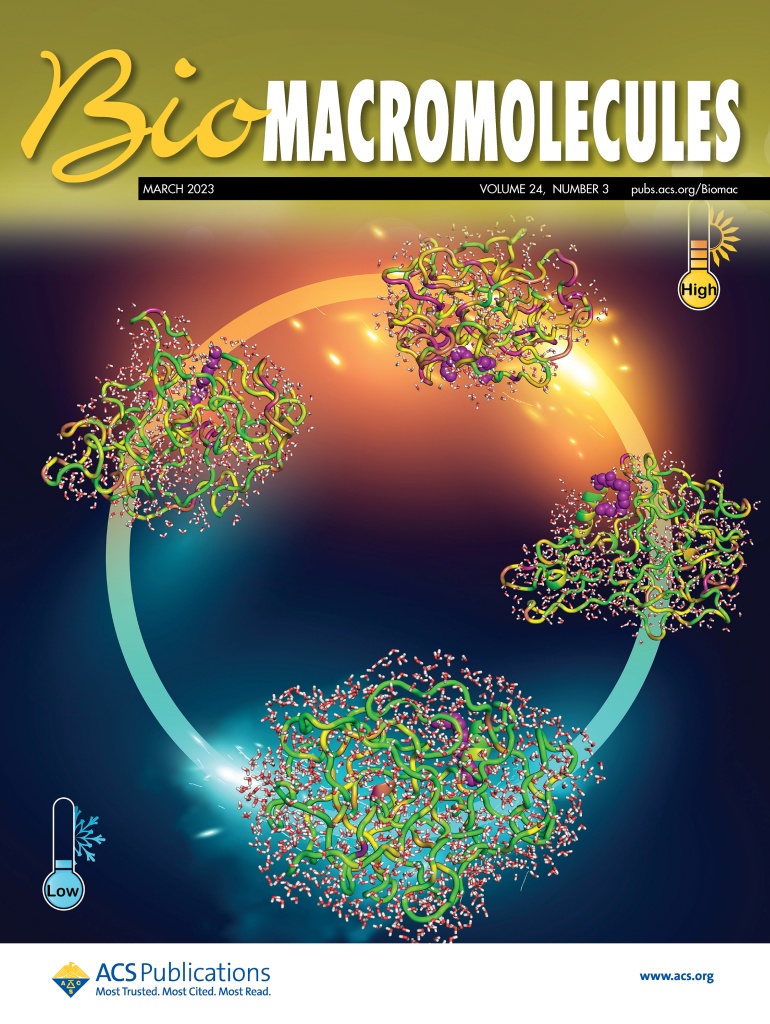
Fat punching above its weight: Modification with lipids yields unexpected alterations in the proteins? structure and hydration.Fatty acid-modified elastin-like polypeptides combine the hierarchical assembly of lipids with the thermoresponsive character of elastins to form nanocarriers with temperature-dependent characteristics. By integrating computational and experimental techniques, we revealed the biophysical underpinnings of thermoresponsive behavior for this class of materials; temperature alters the structure, contact, and hydration of lipid, lipidation site, and protein. These findings enable the rational design of the biomaterials and therapeutics that rival the exquisite hierarchy and capabilities of biological systems
BioMacromolecules 2022
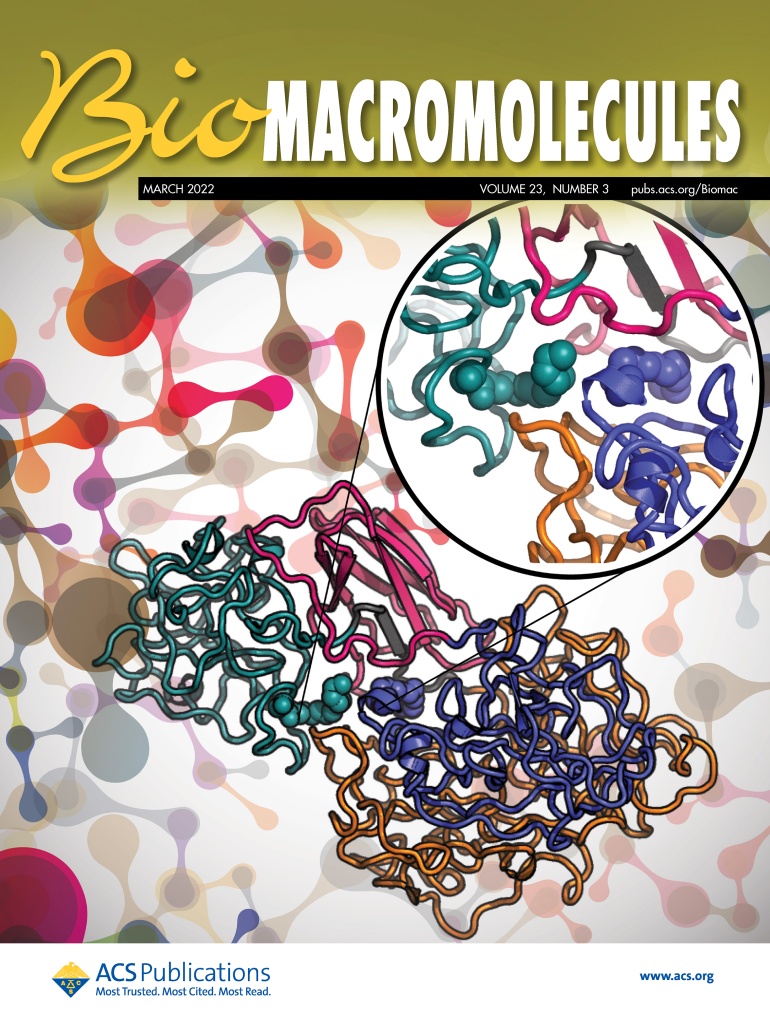
Protein nanoworms are promising candidates for nanomedicine, biotechnology, and synthetic biology, but their promise is constrained by their narrow stability regimes. By integrating experiments, simulations, and data analytics, this study presents a new class of recombinant amphiphiles, post-translationally modified star-shaped lipoproteins, that form adaptive nanoworms with low polydispersity over a broad operational window and reveal their molecular design principles. This integrated approach can accelerate the design and discovery of new nano-biomaterials by reducing the experimental and simulation costs required to identify hybrid biomaterials with desired properties.
Langmuir 2020
Ion transport through the tight junction pores formed by transmembrane claudin proteins between membranes of two adjacent cells. The ion conductance is regulated by the electrostatic environment created by the pore-lining amino acid residues.
Physical Chemistry Chemical Physics 2020
This work investigates the photoactivity of
amphiphilic-coumarin fluorophores in liposomes. We discovered that the liposomal permeability of the coumarin molecule depends on the length of its conjugated-alkyl chain. The short-chain coumarin molecules penetrate the liposomes less effectively than the long-chain molecules. The longer alkyl chains interact favorably with the
hydrophobic lipids of the liposomal bilayer, and the
molecules undergo flip-flops between the two leaflets. The efficacy of long-chain coumarins in liposomes demonstrates the promise of these amphiphilic molecules as fluorophores in cancer detection.
Journal of Chemical Theory and Computation 2019
The protein association energy landscape (PANEL) method provides an extensive data set of membrane protein association in lipid milieu using a combination of stochastic sampling and equilibration simulations. The results are obtained in a fraction of the simulation cost compared to self-assembly simulations.
The Journal of Physical Chemistry B 2018
Molecular architecture of the claudin tight junction proteins. The claudin family of transmembrane proteins forms tight junctions between the adjacent cells in epithelial and endothelial tissues. These tight junctions control the transport and permeability of ions and small molecules through the paracellular space. The tight junction permeability is critical in understanding physiological processes such as the blood–brain barrier, renal filtration, ionic hemostasis, and drug transport across tissue barriers. This multiscale dynamics study of classic claudins provides the molecular architecture of the tight junction pores as well as the putative conformation of claudin proteins that acts as a receptor for the Clostridium perfringens enterotoxin.
Langmuir 2018
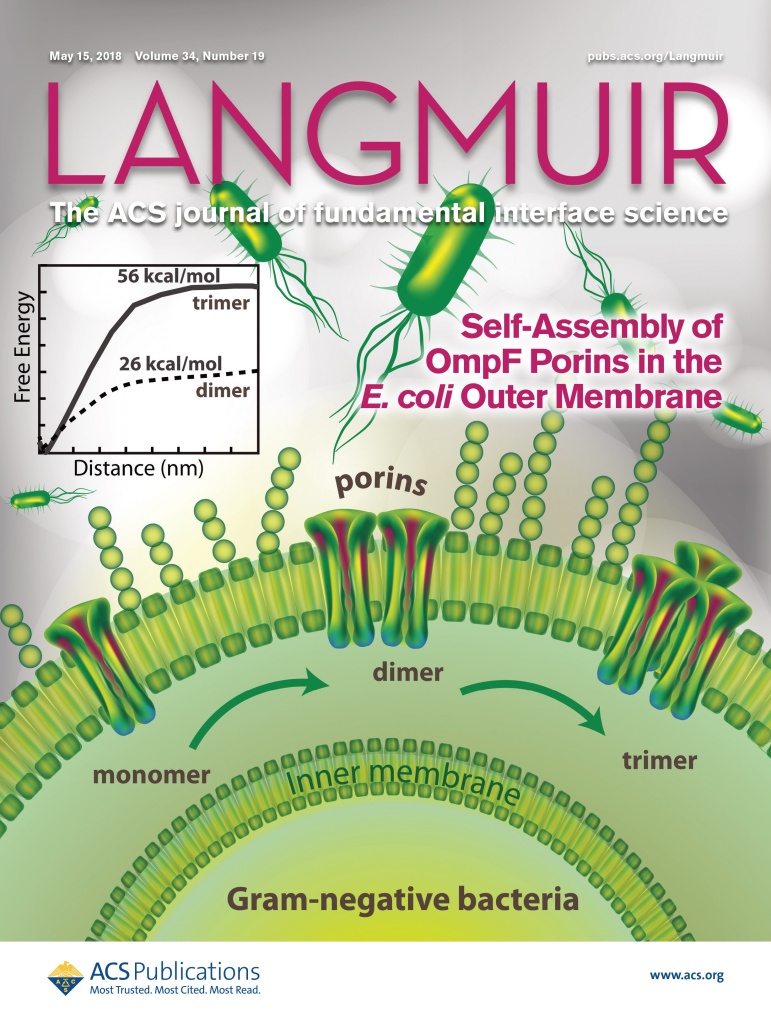
The channel-forming protein OmpF porin from Escherichia coli spans the bacterial outer membrane and facilitates the diffusion-mediated influx of nutrients into the cell. Using molecular dynamics approach, we explore the thermodynamics of OmpF assembly over microsecond timescale simulations, enabled by the newly developed coarse-grain force field parameter set for E. coli outer membrane lipids.
Journal of Chemical Theory and Computation 2015
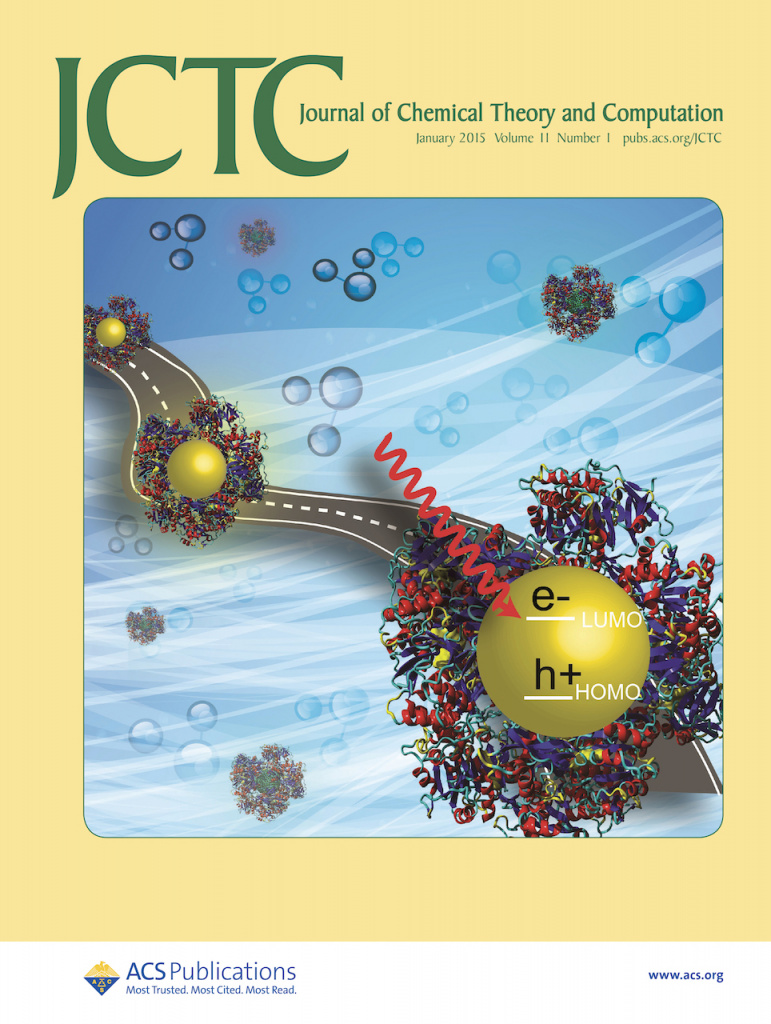
The optical properties of quantum dots are influenced by the presence of surface ligands. Here we investigate the effect of protein corona formation on the optical properties of the firefly Luciferase-CdSe quantum dot complex. To make the computational cost tractable for a large protein-quantum dot system, a multilevel approach was developed where the electronically excited quantum dot was treated quantum mechanically (QM) using electron-hole quasiparticle representation and the protein corona was represented using the molecular mechanics (MM) force field.
The journal of Physical Chemistry B 2015
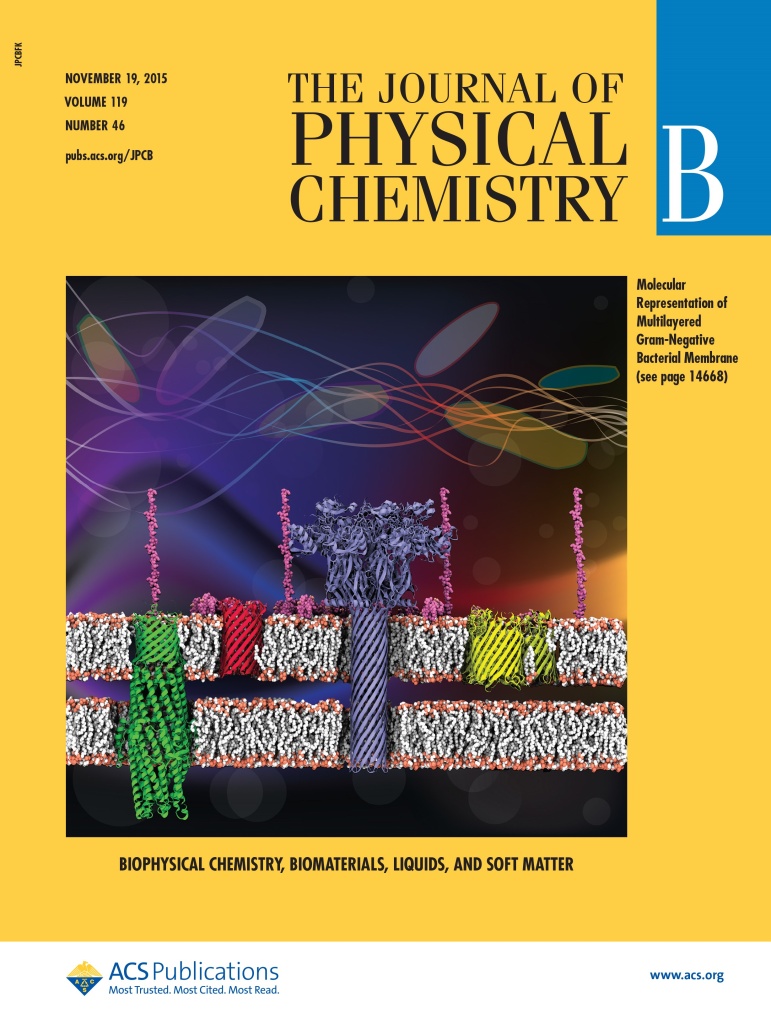
The cell envelope of Gram-negative bacteria is a complex multilayered structure that, besides gatekeeping chemical transport in and out of the bacterial cell, protects the cell from hostile environments. Until now, molecular-level understanding of the bacterial membrane, especially over long time scales, has been challenging due to the enormous computational cost associated with molecular simulations of such complex structures. Here, we have developed and benchmarked a coarse-grained force field for the lipopolysaccharide macromolecule with the O-antigen domain that will enable long timescale molecular simulations of bacterial membranes with fractional computational cost.
Langmuir 2012
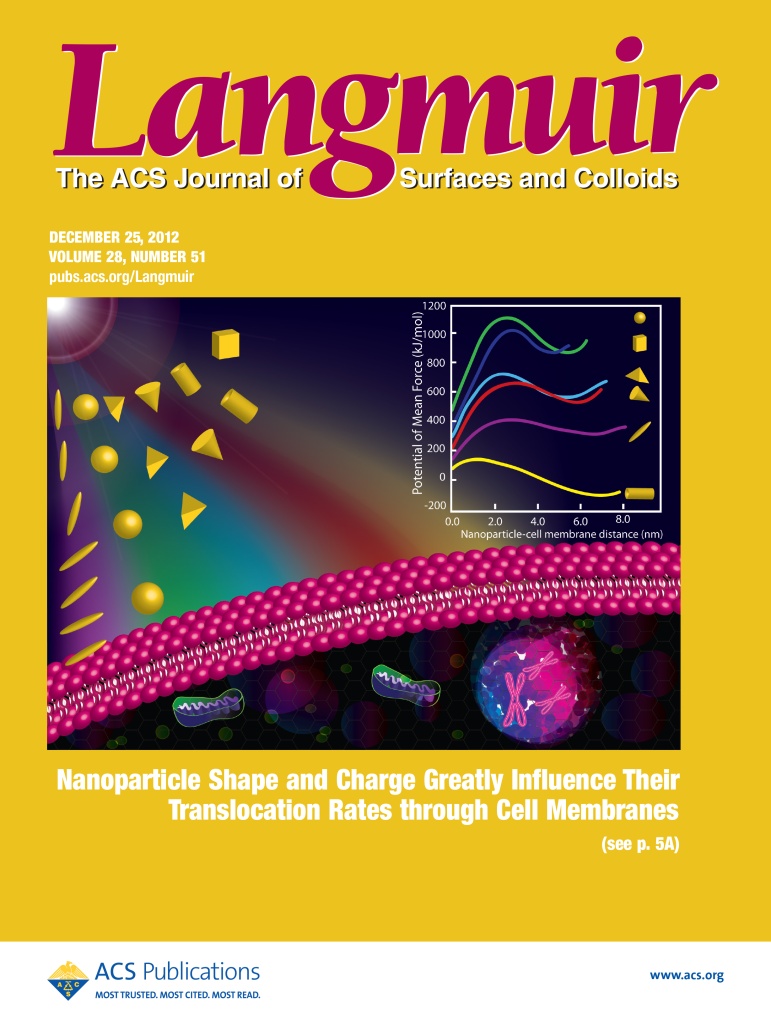
High-performance molecular dynamics simulations were employed to calculate the potential of mean force associated with nanoparticle translocation as well as translocation rate constants. The results indicate that the translocation rate can vary greatly based on the particle surface charge and shape. The simulations also unravel a fascinating electrostatics-driven reorientation mechanism of faceted nanoparticles that facilitates instantaneous transport resulting in the disruption of the self-assembly of the membrane lipids.
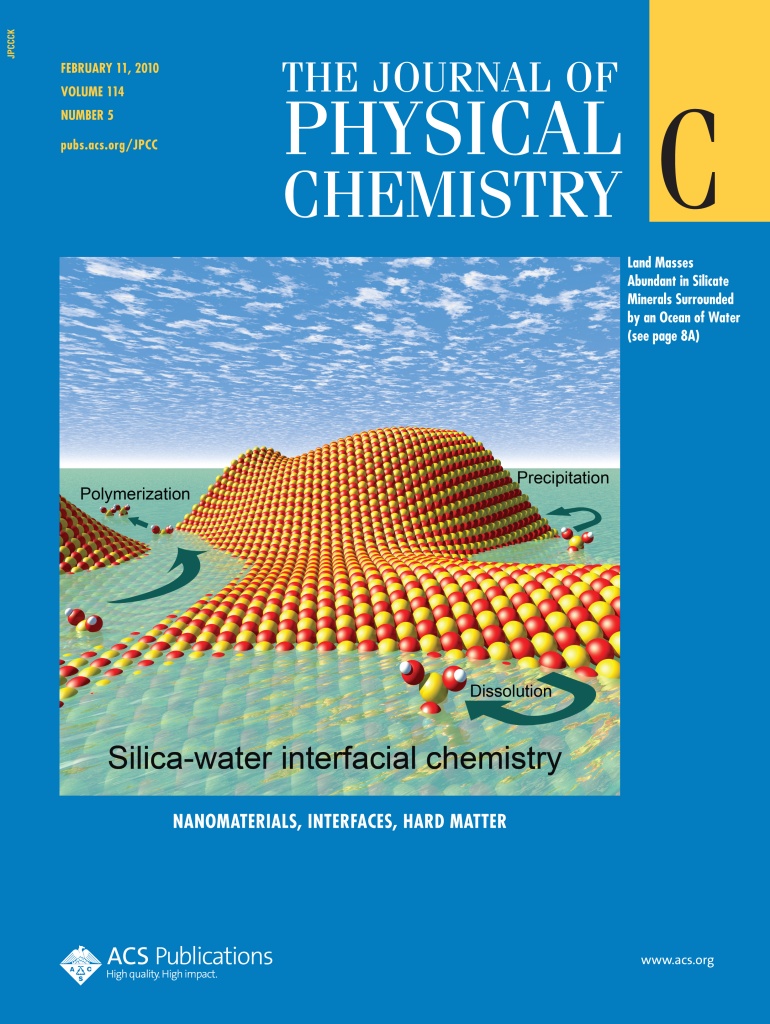
The journal of Physical Chemistry 2010
Land masses abundant in silicate minerals surrounded by an ocean of water. Interfacial silica–water chemistry involves competing dissolution, precipitation, and polymerization reactions. These reactions are ubiquitous on Earth’s surface and occur over geological time scales. A new time-independent advanced Monte Carlo algorithm has been developed that provides insight into reaction mechanisms, equilibrium properties, and surface topography. In the present application of this method, the role of intrasurface hydrogen bonding in the dissolution process is explored.
The Journal of Physical Chemistry 2006
Potential Energy Surfaces for Photodissociation of Ammonia through a Conical Intersection Special Issue honoring Donald G. Truhlar, J. Physical Chemistry A 110, (2006).